Video wird geladen...
Video konnte nicht geladen werden
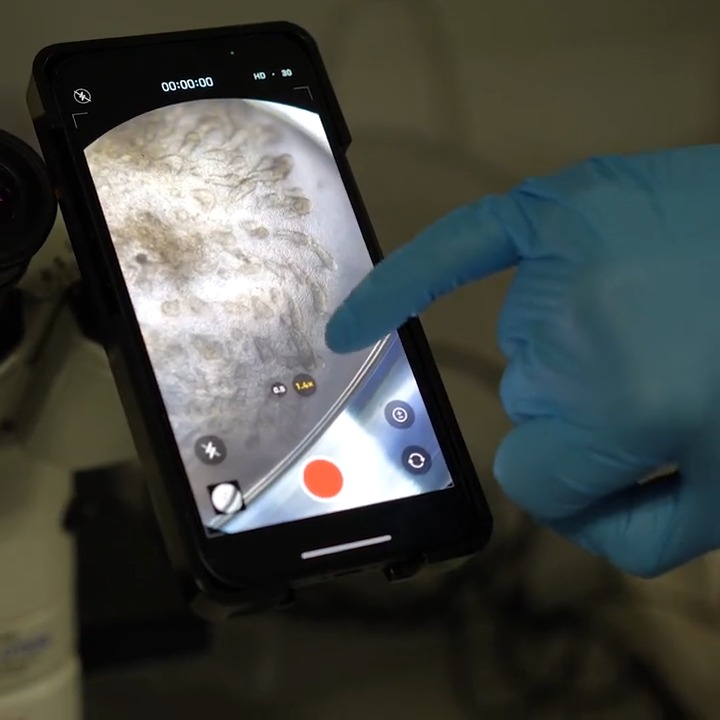
Single-cell technologies now let us profile entire transcriptomes in individual cells. But how do we make sense of this complexity in a biologically meaningful way? Many methods summarise cells into a single embedding, but this often comes at the cost of interpretability, especially when multiple gene programs are active... show more
22,578 Aufrufe • vor 7 Monaten •via X (Twitter)
7 Kommentare

Kavenvor 7 Monaten
AI link for summary and cited qa

Mo Lotfollahivor 7 Monaten
Ahh this is beautiful 😊
Kavenvor 7 Monaten
I love them, you can generate these editable pptx on nouswise for free.

Dr. Zeynep Hülya Gümüş 🕊️vor 7 Monaten
Congrats, Mo! 🔥

Jamie Timmonsvor 7 Monaten
That first line isn't really true. They provide a partial, part biased, part stochastic, SAMPLE of the transcriptome - and the impact of these limitations is as of yet isn't fully understood.

Aiden M Sababivor 7 Monaten
Great work, congrats.

Jaimevor 7 Monaten
@HaoYin20 Clean approach — representing cells through multiple gene programs instead of a single embedding makes a lot of sense. Could the key question be not just which programs are active, but whether they stay dynamically accessible under perturbation?